



Expo
view channel
view channel
view channel
view channel
view channel
view channel
view channel
view channel
view channel
Clinical Chem.Molecular DiagnosticsHematologyImmunology
PathologyTechnologyIndustry
Events
Webinars

- Blood Biomarker Patterns Could Support Monitoring of Anti-Amyloid Treatment
- Prehospital Blood Test Could Reduce Emergency Transfers for Chest Pain
- First-of-Its-Kind Rapid Blood Test Gains CE Mark to Support Prehospital Stroke Triage
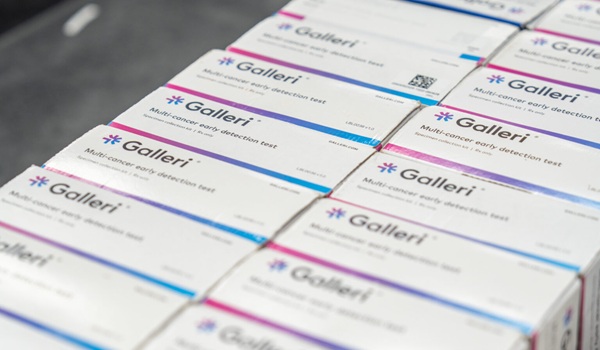
- Multi-Cancer Blood Test Expands Detection to Cancers Lacking Routine Screening Options
- Blood Test Could Guide Drug Selection to Prevent Repeat Heart Attacks and Strokes
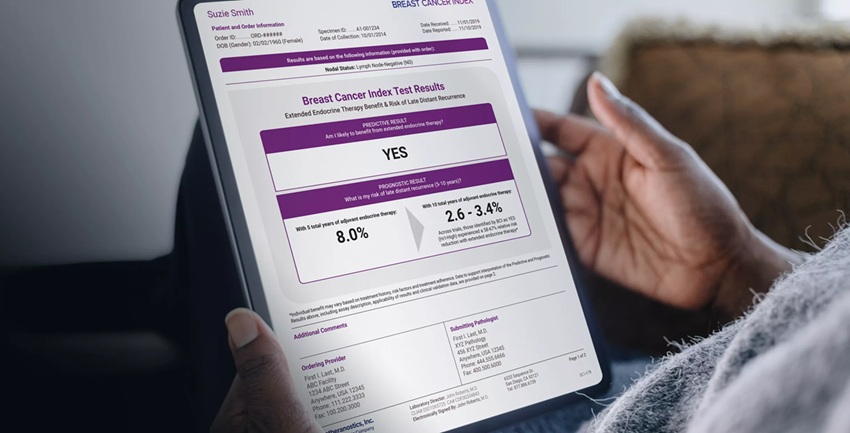
- Genomic Test Guides Extended Endocrine Therapy in Early Breast Cancer
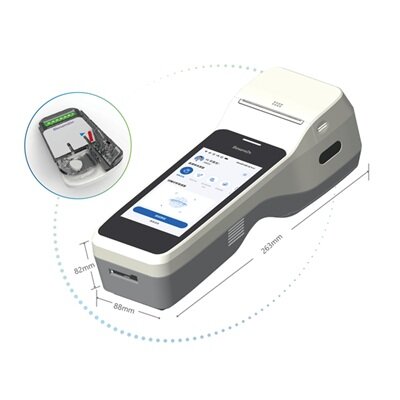
- Portable Test Differentiates Seven Mosquito-Borne Viruses at Point of Care
- Molecular Skin Test May Help Guide Treatment Selection in Atopic Dermatitis
- Genetic Testing Program Helps Uncover Inherited Risk in Pediatric Cancer
- Genomic Breast Cancer Study Adds Real-World Evidence Across Diverse Patient Populations
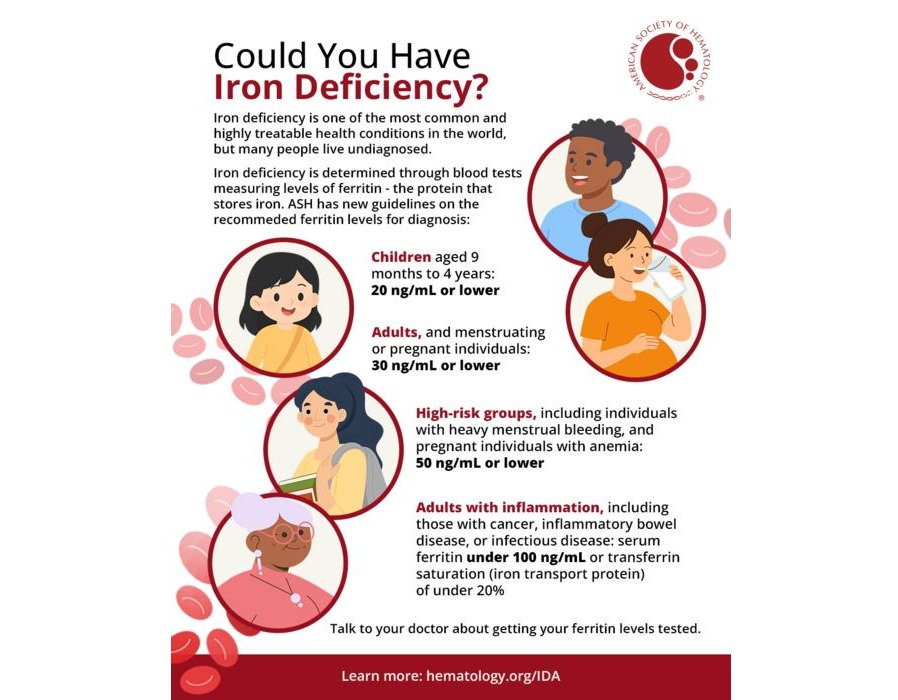
- Updated Ferritin Thresholds Improve Detection of Iron Deficiency
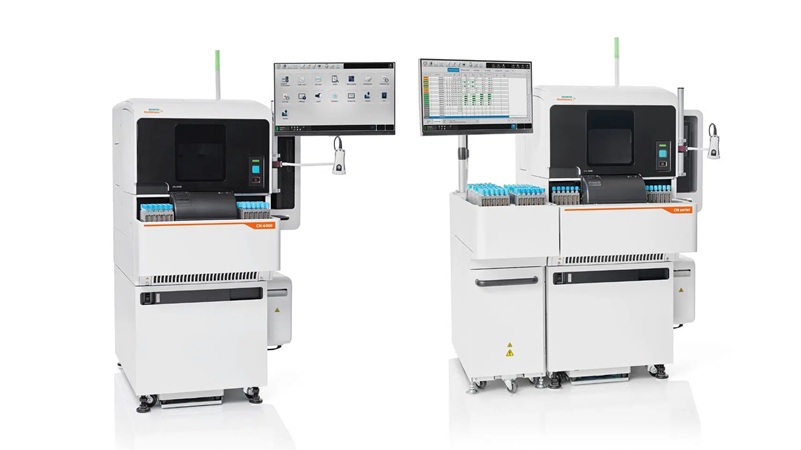
- New Automated Hemostasis Systems Become Available Across US Laboratories
- Variation in AML Residual Disease Testing Prompts Push for Standardization
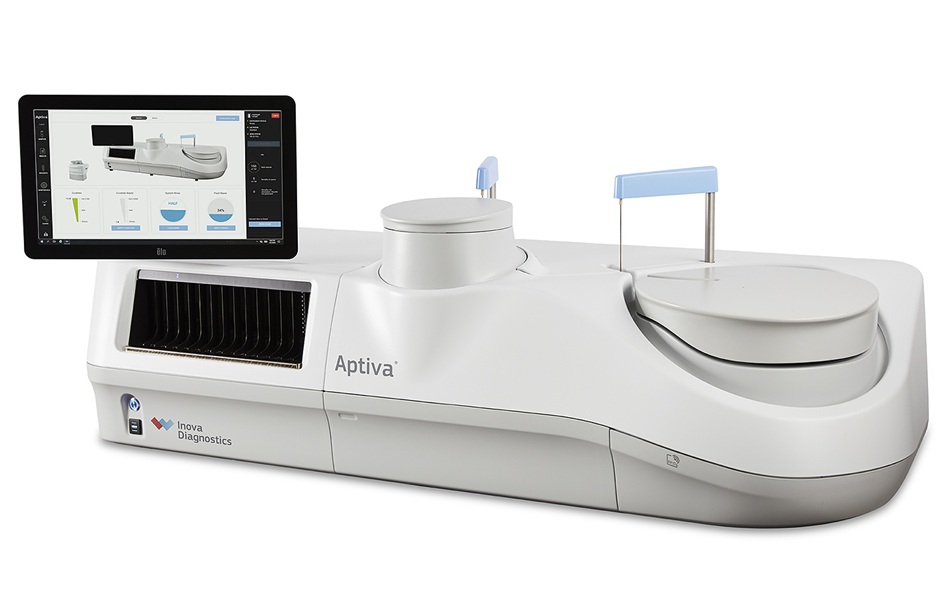
- Hidden Molecular Mechanism Behind HIT Blood-Clotting Disorder Revealed
- Borderline Anemia May Carry Higher Risk Than Current Thresholds Suggest
- Blood Biomarker May Predict Colitis Risk in Checkpoint Inhibitor Therapy
- Blood EBV Activity Biomarkers May Predict Multiple Sclerosis Relapse Months Ahead
- New Cellular Map May Help Predict Crohn’s Disease Course in Children
- Temporal Immune Profiling Reveals How Sepsis States Change Over Time
- Study Identifies Immune Cells That Drive Harmful Autoantibody Responses in COVID-19
- New Urine Test Expands Mycotoxin Analysis to 31 Markers for Broader Exposure Assessment
- Collaboration Targets Blood-Based Ebola Detection Outside Central Laboratories
- Study Finds Frequent Overlap in Antibodies to Lyme and Other Tick-Borne Pathogens
- Early-Life Gut Microbiome Changes May Help Assess Type 1 Diabetes Risk
- Expanded Newborn Screening Program Increases Detection of Congenital Cytomegalovirus
- ADLM Calls for CLIA Updates to Support Safe AI Use in Laboratory Medicine
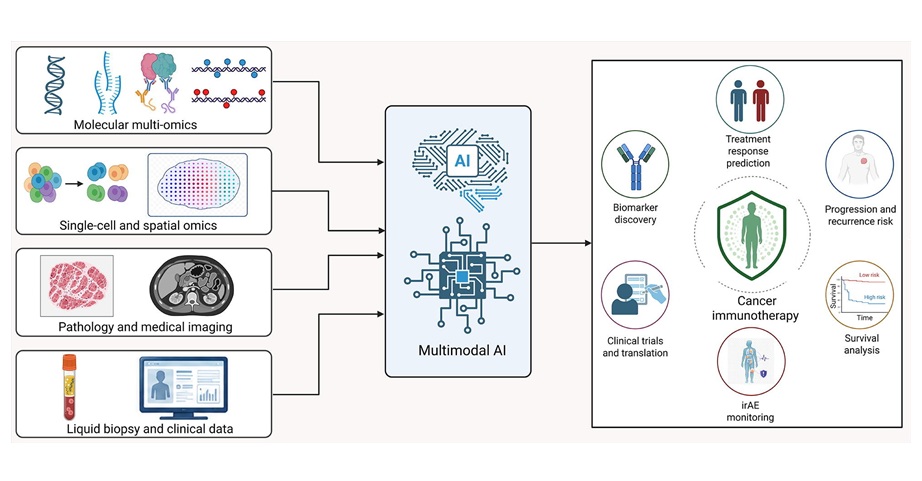
- Multimodal AI Framework Aims to Guide Cancer Immunotherapy Decisions
- New Multipurpose Centrifuge Combines High Capacity with Sustainable Cooling
- Bacterial Vesicle Expression System Streamlines Production of Cancer Diagnostic Proteins
- Autonomous Robotic System Receives FDA Authorization for Blood Collection
- Mayo Clinic Laboratories and Pathology Asia Expand Genomic Testing Across Asia-Pacific
- New UK Collaboration Expands Access to Integrated Alzheimer’s Diagnostic Testing
- Multi-Cancer Early Detection Test Wins FDA Panel Backing for Approval
- Siemens Healthineers and Novo Collaborate to Expand Access to Noninvasive Liver Testing
- Mayo Clinic and Thermo Fisher Launch Multi-Omics Venture to Identify Early Disease Signals
- New Autoantibody Target Identified in Neuromyelitis Optica Spectrum Disorder
- New Gene-Disease Link May Help Diagnose Rare Neurodevelopmental Disorders
- Researchers Identify Shared Molecular Networks Behind Fatigue-Related Illnesses
- New Immune Target Could Support More Precise High Blood Pressure Treatment
- Whole-Blood Extracellular Vesicle Analysis Captures Molecular Signals Missed by Plasma Testing
- Tumor Blood Vessel Features May Help Predict Colorectal Cancer Survival
- Multimodal AI Improves Breast Cancer Recurrence Risk Prediction Beyond Standard Genomic Testing
- Machine Learning Cytology Tool Improves Cancer Cell Identification
- Tumor Budding Grading May Predict Chemotherapy Benefit in Resected Lung Squamous Cancer
- CAP Updates Cancer Protocols to Refine Breast Cancer Biomarker Reporting and GIST Molecular Testing
 Expo
- Blood Biomarker Patterns Could Support Monitoring of Anti-Amyloid Treatment
- Prehospital Blood Test Could Reduce Emergency Transfers for Chest Pain
- First-of-Its-Kind Rapid Blood Test Gains CE Mark to Support Prehospital Stroke Triage
- Multi-Cancer Blood Test Expands Detection to Cancers Lacking Routine Screening Options
- Blood Test Could Guide Drug Selection to Prevent Repeat Heart Attacks and Strokes
- Genomic Test Guides Extended Endocrine Therapy in Early Breast Cancer
- Portable Test Differentiates Seven Mosquito-Borne Viruses at Point of Care
- Molecular Skin Test May Help Guide Treatment Selection in Atopic Dermatitis
- Genetic Testing Program Helps Uncover Inherited Risk in Pediatric Cancer
- Genomic Breast Cancer Study Adds Real-World Evidence Across Diverse Patient Populations
- Updated Ferritin Thresholds Improve Detection of Iron Deficiency
- New Automated Hemostasis Systems Become Available Across US Laboratories
- Variation in AML Residual Disease Testing Prompts Push for Standardization
- Hidden Molecular Mechanism Behind HIT Blood-Clotting Disorder Revealed
- Borderline Anemia May Carry Higher Risk Than Current Thresholds Suggest
- Blood Biomarker May Predict Colitis Risk in Checkpoint Inhibitor Therapy
- Blood EBV Activity Biomarkers May Predict Multiple Sclerosis Relapse Months Ahead
- New Cellular Map May Help Predict Crohn’s Disease Course in Children
- Temporal Immune Profiling Reveals How Sepsis States Change Over Time
- Study Identifies Immune Cells That Drive Harmful Autoantibody Responses in COVID-19
- New Urine Test Expands Mycotoxin Analysis to 31 Markers for Broader Exposure Assessment
- Collaboration Targets Blood-Based Ebola Detection Outside Central Laboratories
- Study Finds Frequent Overlap in Antibodies to Lyme and Other Tick-Borne Pathogens
- Early-Life Gut Microbiome Changes May Help Assess Type 1 Diabetes Risk
- Expanded Newborn Screening Program Increases Detection of Congenital Cytomegalovirus
- ADLM Calls for CLIA Updates to Support Safe AI Use in Laboratory Medicine
- Multimodal AI Framework Aims to Guide Cancer Immunotherapy Decisions
- New Multipurpose Centrifuge Combines High Capacity with Sustainable Cooling
- Bacterial Vesicle Expression System Streamlines Production of Cancer Diagnostic Proteins
- Autonomous Robotic System Receives FDA Authorization for Blood Collection
- Mayo Clinic Laboratories and Pathology Asia Expand Genomic Testing Across Asia-Pacific
- New UK Collaboration Expands Access to Integrated Alzheimer’s Diagnostic Testing
- Multi-Cancer Early Detection Test Wins FDA Panel Backing for Approval
- Siemens Healthineers and Novo Collaborate to Expand Access to Noninvasive Liver Testing
- Mayo Clinic and Thermo Fisher Launch Multi-Omics Venture to Identify Early Disease Signals
- New Autoantibody Target Identified in Neuromyelitis Optica Spectrum Disorder
- New Gene-Disease Link May Help Diagnose Rare Neurodevelopmental Disorders
- Researchers Identify Shared Molecular Networks Behind Fatigue-Related Illnesses
- New Immune Target Could Support More Precise High Blood Pressure Treatment
- Whole-Blood Extracellular Vesicle Analysis Captures Molecular Signals Missed by Plasma Testing
- Tumor Blood Vessel Features May Help Predict Colorectal Cancer Survival
- Multimodal AI Improves Breast Cancer Recurrence Risk Prediction Beyond Standard Genomic Testing
- Machine Learning Cytology Tool Improves Cancer Cell Identification
- Tumor Budding Grading May Predict Chemotherapy Benefit in Resected Lung Squamous Cancer
- CAP Updates Cancer Protocols to Refine Breast Cancer Biomarker Reporting and GIST Molecular Testing

.jpg)



 Client Login